

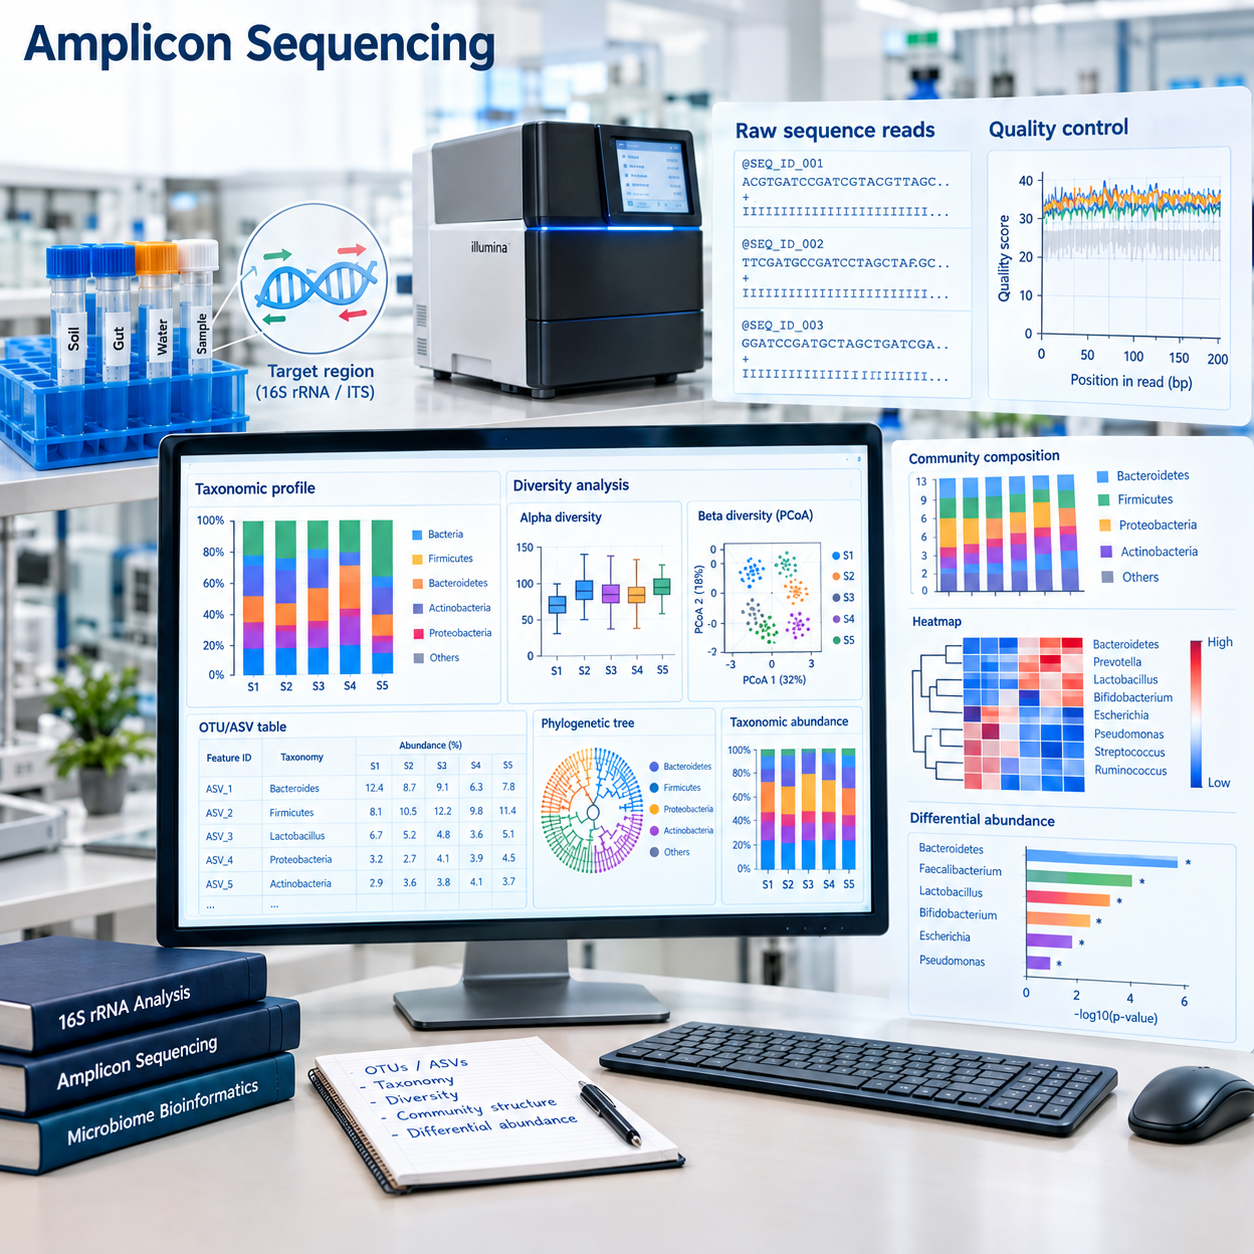
Amplicon Sequencing
Targeted amplicon pipelines for panel-based variant detection, pathogen genotyping or marker screening — built around your primer design, not a generic one-size pipeline.
TYPICAL CLIENT
A molecular diagnostics team running a custom gene panel or pathogen assay
Usually a molecular diagnostics lead or applied genomics scientist running a custom amplicon panel — a cancer hotspot panel, a pathogen genotyping assay or a breeding marker screen — who needs primer-aware analysis, not a pipeline built for whole-genome data.
Six concrete deliverables, not a slide deck.
This is exactly what transfers to you at the end of this Amplicon Sequencing engagement.
Primer-aware demultiplexing & trimming
A trimming step configured to your exact primer scheme, so amplicon boundaries never get miscalled as variants.
Low-frequency variant calling
Variant calling tuned for the low allele fractions common in hotspot panels and pathogen quasispecies.
UMI & duplicate handling
Unique molecular identifier consensus calling where UMIs are used, to separate true low-frequency signal from PCR error.
Panel-specific QC dashboard
Per-amplicon coverage and dropout reporting, so failed amplicons are visible before results are reported out.
Batch-level QC report
Cross-sample QC summarizing coverage uniformity and failure rates across a full sequencing batch.
Full documentation & handoff session
Written documentation plus a recorded walkthrough so your team can operate the pipeline independently.
What this service includes
Primer trimming & demultiplexing
Reads trimmed against your exact primer scheme so amplicon edges are never mistaken for real variants.
Low-VAF variant calling
Calling tuned to detect variants at allele fractions standard germline callers would filter out.
UMI consensus calling
Where UMIs are used, PCR duplicates are collapsed into a consensus read before variant calling.
Per-amplicon panel QC
Coverage and dropout reported at the individual amplicon level, not just averaged across the panel.
What this looks like in a real engagement
Three concrete examples of the kind of work this service covers.
Cancer hotspot panel
A targeted oncology panel analyzed for low-frequency somatic hotspot mutations across dozens of amplicons.
Viral genotyping amplicon panel
A pathogen surveillance panel processed to call lineage-defining variants from patient or wastewater samples.
Agrigenomics marker screening
A breeding program’s marker panel genotyped across hundreds of samples for trait-associated variants.
Why amplicon data needs its own pipeline, not a shrunk-down WGS one.
Amplicon data breaks assumptions a whole-genome pipeline relies on — primer sequences show up at every read boundary, coverage is deliberately non-uniform, and the variants that matter are often at allele fractions a standard germline caller would filter out as noise.
We build amplicon pipelines that trim exactly to your primer scheme and call variants at the sensitivity your panel actually needs, so a low-frequency signal doesn’t get lost in default filtering thresholds.
How an engagement runs
Four stages, from sample intake to delivery.
Demultiplex & primer trim
Reads are demultiplexed and trimmed against your exact primer scheme before any calling starts.
Low-frequency variant calling
Variants are called with sensitivity tuned to your panel’s expected allele fraction range.
Per-amplicon QC
Coverage and dropout are reported per amplicon, flagging any that failed before reporting.
Report & hand off
Batch-level QC and a final variant report are delivered alongside full documentation.
What we build it with
The variant that matters in an amplicon panel is usually the one a default germline caller would throw away as noise.
Before you reach out
With UMI-based panels we routinely detect variants down to 0.5–1% allele fraction; without UMIs, reliable detection is typically 2–5% depending on depth.
Yes — the primer scheme is what makes amplicon trimming accurate, so we’ll ask for it (or help you generate one) at project kickoff.
Yes, including minority variant calling for viral quasispecies, with panel-specific thresholds set for your assay.
Explore related work
Whole Exome Sequencing
Step up to full coding-region coverage when a fixed panel is no longer enough.
View CHIPChIP Sequencing
A different targeted approach — profiling protein-DNA binding instead of genotype.
View BIOBioinformatics Pipeline overview
See how amplicon sequencing fits into the full Bioinformatics Pipeline service.
ViewRunning a panel that needs primer-aware analysis?
Send us your primer scheme and target variants — we’ll tell you honestly what sensitivity your panel design can support.